Revisiting a Journal Club Article, "Restored clearance of senescent neutrophils by tissue-resident macrophages limits organ aging." Science. 2026
Why This Paper Matters
Most of the senescence conversation over the past decade has centered on a familiar cast: senescent fibroblasts, endothelial cells, adipocytes, and the drugs designed to clear them. This paper, which I discuss in my September Journal Club, argues that we have been watching the wrong cell and, more importantly, the wrong failure point.
The authors of the study ask us to consider that organ aging is driven less by senescent cells accumulating on their own and more by the collapse of the system responsible for removing them. As the Stanford group frames it, aging looks less like passive degeneration and more like a failure of active cellular clearance. I think this is a meaningful reframing, because a clearance defect is something we can potentially restore.
Background: Two Cell Types, One Relationship
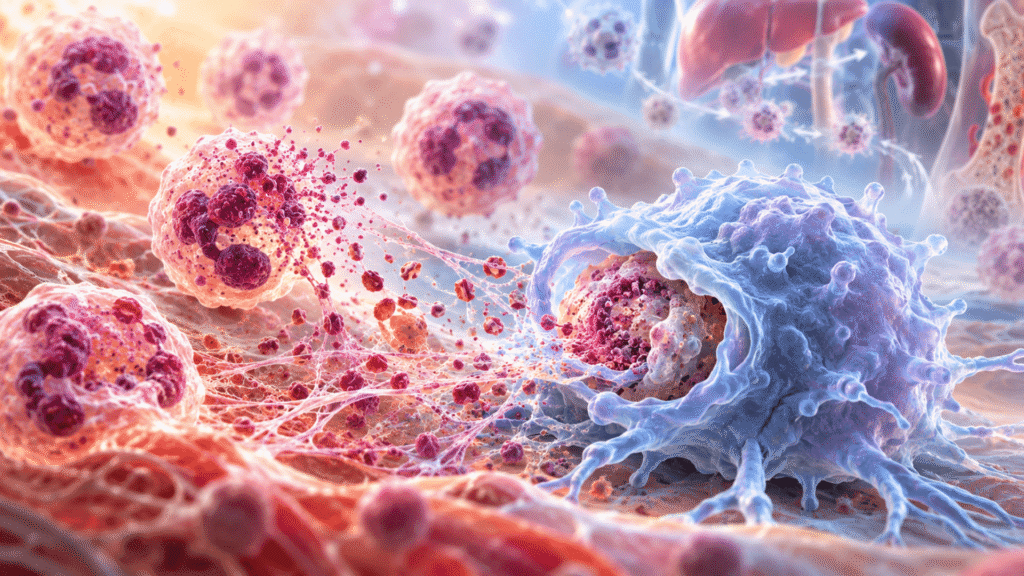
Neutrophils are the most abundantly produced immune cell in the body — on the order of 100 billion generated daily in humans, roughly 10 billion in mice — and among the shortest-lived, surviving only about 12 to 24 hours in circulation. Significantly, neutrophils begin showing senescent markers within 8 to 12 hours of entering the bloodstream, and yet the overwhelming majority never encounter a pathogen: They simply age out, with roughly 90% of them ending up in the liver, spleen, and bone marrow awaiting disposal.
However, when they are not disposed of promptly, they do damage. Senescent neutrophils degranulate, release proteases, and undergo NETosis — the extrusion of neutrophil extracellular traps. Those products break down lipid membranes and extracellular matrix, generate oxidative stress and DNA damage in bystander cells, and release cytokines and chemokines that recruit still more neutrophils into the tissue. This creates a powerful self-reinforcing loop.
Tissue-resident macrophages (TRMs) are the counterweight to this cycle. They take up residence in organs during fetal development and stay for the life of the animal, adapting to the tissue they inhabit. They make up roughly 60 to 90% of the macrophage population in the brain, liver, lung, heart, and kidney. Their core job is efferocytosis: recognizing, engulfing, and digesting apoptotic, damaged, and senescent cells before those cells become an inflammatory liability. However, this long lifespan is also their vulnerability. Decades of accumulated metabolic, oxidative, and inflammatory injury degrade their function.
The Mechanism: EP2
The link between the neutrophils and TRMs is prostaglandin E2 and its receptor EP2. PGE2 output rises with age, with infection, with injury, and with toxic exposure. EP2 is the pro-inflammatory member of the PGE2 receptor family, and tissue-resident macrophages are densely loaded with it. EP2 expression itself also increases with age.
EP2 triggers a specific chain reaction inside cells: it turns on a protein called adenylyl cyclase, which boosts cyclic AMP (cAMP) levels, and ultimately activates protein kinase A (PKA). PKA then acts on transcription factors in the nucleus that govern gene expression, including the expression of integrins on the macrophage surface. Those integrins are the recognition apparatus that enable the macrophage to engage a senescent neutrophil and initiate engulfment. As the authors of the study indicate, EP2 signaling suppresses macrophage metabolism and phagocytosis; in addition, this rising EP2 activity shifts the transcriptional program so that integrin-mediated recognition and phagolysosomal engulfment both fail. As a result, the neutrophil is left behind to degranulate and NETose, and its senescence-associated secretory phenotype (SASP) does its damage unchecked. Removing or blocking EP2 restores the interaction and the engulfment, which in turn limits degranulation, NETosis, paracrine stress, and organ aging, the causal chain laid out in the paper's own summary figure.
Study Design and Results
The work was done in male mice, with a human validation arm. The investigators built a conditional knockout (Cx3cr1CreER; Ptger2lox/lox), which allowed them to delete the EP2-encoding gene Ptger2 specifically in tissue-resident macrophages at a chosen time. Tamoxifen was given at 4 to 6 months of age, the murine equivalent of adolescence. A ubiquitous conditional knockout line and littermate wild-type controls were run alongside. Young mice at 6 to 8 months (human late adolescence/early adulthood) were compared against aged mice at 23 to 25 months (human 60s to 70s), with and without TRM EP2 deletion. In addition, a large single-cell atlas of young, aged, and diseased human liver was interrogated for the same signature.
It's worth noting that the study was limited by only studying male mice. Given that PGE2 signaling, inflammatory tone, and neutrophil biology all show sex differences, whether these results generalize to females is an open question the study does not answer.
In aged control mice, senescent neutrophils built up most heavily in liver, spleen, and bone marrow, with lesser accumulation across other organs. In aged mice without TRM EP2, neutrophil numbers stayed near youthful levels. The EP2-deleted aged mice were leaner, carried less visceral fat, retained more muscle mass, and performed on multiple organ-function tests at the level of young animals. Speed, balance, and grip strength resembled those of young mice. Inflammation was reduced in blood, liver, colon, heart, kidney, and hippocampus. Memory was preserved — maze navigation and object recognition were near-young and far better than aged controls with functional EP2. Mitochondrial fitness was maintained.
Two months of PF-04418948 in 22-month-old mice reduced both total and senescent neutrophil counts toward youthful values. In culture, aged TRMs showed impaired capacity to engulf and digest spent neutrophils; the EP2 antagonist significantly restored it. The rescue was strongest in the liver. This is the key translational result: the defect is reversible late in life, not merely preventable early.
The human liver atlas revealed age-related neutrophil accumulation, increased neutrophil senescence, TRM decline, and elevated EP2 activity — more pronounced still in diseased livers. The authors are explicit that these findings are correlative and require functional testing.
My Key Takeaways
First, I think it’s important to consider that the senescent neutrophil is under-appreciated. As I’ve discussed in several of my books, senescence occurs in many cell types, but this paper isolates one we have largely ignored and shows that senescence in this single population is enough to drive tissue damage and organ-wide aging. Second, it’s also significant that the defect is transcriptional, and it lives in the macrophage. The lesion is not primarily in the senescent cell; it’s in the transcriptional programming of the macrophage that is supposed to remove it:
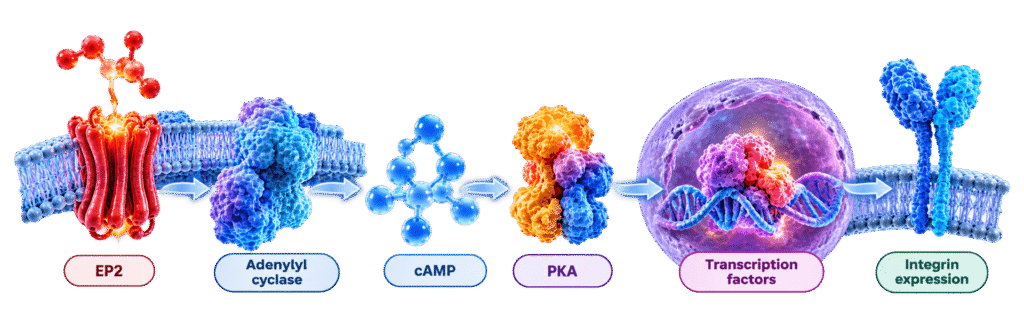
Integrins are adhesion and signaling molecules central to cell-to-cell interaction, and when their expression is disrupted, recognition fails.
Third, signaling tone matters more than presence or absence. A normal macrophage requires some EP2 signaling to drive the transcription factors that regulate integrin communication. The pathology reflects an excess, i.e., too much EP2 activity, or abnormalities in the receptor. This is a dose problem, not an on/off switch, which has direct implications for how any intervention should be designed.
Fourth, keep in mind that efferocytosis is a reparative process that should be protected. When a cell undergoes apoptosis, membrane breakdown externalizes phosphatidylserine, which serves as the "eat me" signal. The macrophage engulfs the cell, degrades it through the lysosomal pathway, and repurposes the membrane components. Efferocytosis is anti-inflammatory and restorative by design; it’s not simply garbage removal.
I also think it’s important to understand that the loop is self-amplifying, and stress feeds it. Failed clearance means more protease release and NETosis, more membrane and tissue damage, more oxidative stress and DNA damage, and more chemokine-driven neutrophil recruitment. Compounding this, inflammatory cytokines (TNF-α and IL-6 among them) upregulate EP2 itself. The stress state worsens the very receptor abnormality that caused the failure.
Finally, the study targets the receptor, but PGE2 is generated by COX-1 and COX-2 acting on arachidonic acid, and COX activity and arachidonic-acid-driven PGE2 production both rise with aging. If PGE2 is the bad player, the upstream question is where that PGE2 is coming from. That is the real question this paper points toward: what is the source and driver of EP2 activation?
The specialized pro-resolving mediators that govern the resolution phase of inflammation sit directly in this pathway. Ceramides may serve as a useful indicator of how the process is progressing. And genetic contributors — SNPs in the EP2 receptor gene and their relationship to aging — deserve investigation.
Keep in mind, too, that this is a mechanistic framework, not a product. There are no approved drugs that selectively block EP2. NSAIDs act upstream on PGE2 production but block other prostaglandins with legitimate functions, and PGE2 itself is beneficial through receptors other than EP2. The value here is that a previously undescribed failure point has been isolated, one that connects to upstream mechanisms many of us are already working to influence.
My bottom line? A single receptor on a single long-lived cell type governs whether the body clears the 100 billion neutrophils it produces every day. When that clearance fails, the debris ages the organ. When it is restored — genetically or pharmacologically, even in old animals — measures of organ function move back toward youthful. The findings are preclinical, conducted in male mice only, and the human data are correlative. But the mechanism is now on the table, and the upstream question is the one worth chasing.
Sources
Tan, Y. J., Conley, T. E., Yao, F., García-Marqués, F. J., Akinyemi, D. E., Dinh, V. V., Wang, Q., Bermudez, A., Kim, J., Belk, J. A., Soehnlein, O., Pitteri, S. J., & Andreasson, K. I. (2026). Restored clearance of senescent neutrophils by tissue-resident macrophages limits organ aging. Science (New York, N.Y.), 393(6808), eaea3075. https://doi.org/10.1126/science.aea3075